skip to content

How to Choose a Biopharma Manufacturing Partner?

Choosing a biopharma manufacturing partner is not a simple vendor comparison. The decision can shape product quality, supply continuity, patient safety, and launch timing. A capable partner should match your molecule, process, scale, and development stage. A persuasive website is not enough. Walk through the facility. Review how operators document deviations, control materials, and protect data. Ask who will manage your program after the contract is signed. Small details matter. A mislabeled sample or unclear handoff can create expensive delays.

This guide examines the evidence behind a reliable Biopharma Manufacturing partnership. It considers technical platforms, quality systems, regulatory readiness, capacity, technology transfer, and communication. Request recent audit findings, inspection history, validation summaries, and realistic production timelines. Confirm whether the site has experience with your formulation, aseptic operations, analytical methods, or cold-chain needs. Financial stability also deserves attention, especially for long development programs. Experts should explain risks plainly, not hide them behind polished language. No partner is flawless. The real question is how quickly and transparently problems are identified, contained, investigated, and corrected.

A strong evaluation also measures cultural fit. Your team may need rapid decisions during a failed batch, changing specification, or shipment disruption. Define escalation routes, decision rights, reporting frequency, and success metrics before work begins. Speak with current or former clients when possible, while recognizing that references can be selective. Independent quality and regulatory advice can test optimistic assumptions. The best choice is rarely the cheapest quote. It is the partner that can repeatedly deliver compliant, traceable, and scientifically justified work. Even then, review the relationship regularly. Conditions change, and early confidence can become complacency.
How to Choose a Biopharma Manufacturing Partner?

Defining Your Biopharma Manufacturing Requirements

How to Choose a Biopharma Manufacturing Partner?

Defining Your Biopharma Manufacturing Requirements

A suitable manufacturing partner begins with a precise requirement sheet. Define the molecule, expression system, dosage form, batch size, and development stage. Specify critical quality attributes, analytical methods, and acceptable process ranges. Include temperature limits, shipping conditions, and planned commercial markets. Small omissions become expensive during technology transfer.

The FDA’s Drug Shortages Task Force reported that manufacturing and quality problems contributed to 62% of drug shortages. This finding makes quality systems a core selection requirement, not a compliance formality. Request recent inspection outcomes, deviation trends, batch-release timelines, and corrective-action evidence. Ask how the partner manages raw-material changes and supplier failures. Their answers should be measurable.

BioPlan Associates’ 2024 biomanufacturing survey collected feedback from more than 300 industry professionals. Its continuing focus on capacity, outsourcing, and production constraints reflects a practical concern: available space does not always mean available capability. Confirm reserved suites, qualified equipment, trained operators, and realistic scheduling. Review process-development support, scale-up data, and comparability experience. A polished presentation can still hide weak execution.

Be specific about governance. Name decision owners, meeting frequency, escalation routes, and documentation standards. Define success for each transfer milestone. I would also test the uncomfortable assumptions. Is the projected yield based on real batches? Can the partner support your peak demand? What happens if the timeline slips? A perfect plan rarely survives contact with production.

How to Choose a Biopharma Manufacturing Partner? — Defining Your Biopharma Manufacturing Requirements

This planning scorecard allocates 100% of the partner-selection focus across the requirements most commonly assessed in biopharmaceutical manufacturing: GMP and regulatory readiness, quality systems, technical capability, capacity, supply-chain resilience, and technology transfer. The weighting is a practical evaluation framework rather than company-specific performance data.

Evaluating Manufacturing Capabilities and Technical Expertise

Choosing a biopharma manufacturing partner requires more than reviewing equipment lists. Technical capability appears in daily execution, especially during scale-up, process transfer, and unexpected deviations.

Examine whether the facility can support your product’s full development stage. Review batch sizes, equipment flexibility, environmental controls, and analytical testing capacity. Ask how engineers transfer a process from laboratory vessels to production tanks. The answer should include defined parameters, risk assessments, comparability studies, and documented training.

Request evidence, not polished promises. Study anonymized batch records, validation protocols, deviation reports, and corrective action plans. A mature partner can explain recurring problems without hiding them. That matters.

Technical expertise also depends on people. Meet process scientists, manufacturing supervisors, quality specialists, and analytical leads. Ask who will make decisions during a failed assay or unexpected yield change. Their responses reveal practical experience better than a presentation.

Data integrity should be visible in routine work. Check electronic systems, audit trails, sample controls, and review procedures. Confirm that analytical methods are suitable for the product and verified at the intended site. Small gaps can become expensive delays.

Facility tours can be revealing. Look for clear material flows, organized production areas, calibrated instruments, and realistic maintenance schedules. A spotless room is not enough. I have found that conversations with operators often reveal more than formal meetings.

No partner is perfect. The stronger one admits limitations and proposes measurable improvements. You may still miss something during evaluation. Build technical agreements with clear responsibilities, escalation routes, change controls, and performance metrics before the first engineering batch.

Assessing Quality Systems, Compliance, and Regulatory Readiness

How to Choose a Biopharma Manufacturing Partner?

A reliable partner makes quality visible, not merely attractive during a sales presentation. Review the quality management system, deviation records, CAPA effectiveness, and batch-release trends. Ask to see anonymized examples. Check whether investigations identify root causes or simply assign operator error. The FDA’s Drug Shortages Task Force report found that quality problems contributed to 62% of drug shortages. In 2023, the FDA reported that manufacturers and regulators helped prevent 236 shortages. These figures show why quality maturity directly affects supply continuity.

Regulatory readiness requires more than a clean inspection history. Examine data-integrity controls, electronic audit trails, change-control discipline, and validation evidence. Confirm that the site can manage technology transfer without losing critical process knowledge. Review inspection responses from the past three years. Speak with quality leaders, not only business development staff. I would not treat a perfect audit score as proof of control. A polished tour can still hide weak documentation. The better question is whether problems are reported early and corrected permanently.

Tips: Request a mock inspection with independent reviewers. Test one deviation from event to CAPA closure. Check training records at the production floor. Compare written procedures with actual operator behavior. Confirm regulatory timelines for your target markets. The FDA’s 2023 drug-shortage report and ICH Q10 principles provide useful reference points, but local requirements still need careful interpretation.

How to Choose a Biopharma Manufacturing Partner? - Assessing Quality Systems, Compliance, and Regulatory Readiness

Assessment Dimension What to Verify Relevant Requirement or Benchmark Objective Evidence to Request Suggested Weight Score
(1–5)
Weighted Result
Pharmaceutical Quality System Assess whether the quality system is formally established, maintained, and independently overseen. ICH Q10 principles, including management responsibility, process performance, product quality monitoring, CAPA, change management, and management review. Quality manual, organization chart, quality objectives, management-review records, annual product-quality reviews, and recent quality metrics. 15% — —
GMP Compliance History Review inspection outcomes, recurring observations, response quality, and remediation effectiveness. Applicable GMP requirements, including 21 CFR Parts 210 and 211 for finished pharmaceuticals and relevant EU GMP requirements. Inspection reports, warning or deficiency correspondence where applicable, CAPA plans, effectiveness checks, and inspection-readiness procedures. 15% — —
Data Integrity and Computerized Systems Determine whether electronic and paper records are complete, attributable, legible, contemporaneous, original, accurate, consistent, enduring, and available. ALCOA+ data-integrity principles, 21 CFR Part 11 where applicable, and EU GMP Annex 11 expectations. Data-integrity risk assessments, audit-trail reviews, user-access reviews, backup and disaster-recovery tests, system-validation reports, and electronic-record procedures. 12% — —
Contamination Control and Aseptic Capability Evaluate facility design, environmental monitoring, personnel practices, cleaning, sterilization, and contamination-control strategy. EU GMP Annex 1 for sterile medicinal products, including contamination-control strategy and ongoing environmental monitoring. Facility layouts, classified-area data, environmental-monitoring trends, media-fill results, smoke studies, cleaning-validation reports, and deviation history. 12% — —
Process Validation and Continued Process Verification Confirm that critical process parameters and critical quality attributes are scientifically understood and controlled throughout the lifecycle. FDA process-validation lifecycle approach: process design, process qualification, and continued process verification; supported by ICH Q8 and Q9 principles. Process-development reports, risk assessments, qualification protocols, validation reports, control strategy, statistical trend reports, and annual review data. 12% — —
Analytical and Laboratory Controls Assess method suitability, laboratory controls, reference standards, stability testing, and out-of-specification investigation practices. GMP laboratory controls, validated or appropriately qualified analytical methods, stability expectations, and scientifically sound OOS investigations. Method-validation or verification protocols, system-suitability data, stability protocols, OOS/OOT procedures, laboratory error investigations, and analyst-training records. 10% — —
Deviation, CAPA, and Change Control Check whether issues are investigated using documented root-cause methods and whether changes are risk-assessed before implementation. ICH Q9 quality-risk-management principles and ICH Q10 CAPA and change-management expectations. Aged-deviation reports, CAPA effectiveness checks, change-control records, risk assessments, escalation rules, and recurring-event trend analyses. 10% — —
Regulatory Submission and Inspection Readiness Evaluate the ability to provide accurate, traceable, and timely information for regulatory submissions, questions, and inspections. Documented regulatory-information governance, controlled responses, product-quality knowledge, and inspection-management procedures. Mock-inspection results, response-management procedures, submission-support experience, document-retrieval tests, regulatory commitments, and contact escalation paths. 8% — —
Supplier and Material Control Review qualification, monitoring, testing, traceability, and change-notification controls for critical materials and outsourced services. Risk-based supplier qualification, approved-supplier controls, material traceability, and quality agreements for critical activities. Supplier-audit reports, qualification procedures, material specifications, certificates of analysis, supplier-performance metrics, and notification commitments. 8% — —
Quality Agreement and Governance Confirm clear allocation of responsibilities for batch release, deviations, complaints, recalls, changes, audits, and regulatory communication. Written quality agreement aligned with applicable GMP responsibilities and the product’s regulatory filing strategy. Draft quality agreement, responsibility matrix, governance calendar, escalation timelines, communication records, and business-continuity procedures. 5% — —
Total 100% — —
Scoring Guide: 1 = unacceptable or no objective evidence; 2 = significant gaps; 3 = meets minimum expectations; 4 = robust and consistently demonstrated; 5 = mature, proactive, and independently verified.
Decision Rule: Treat unresolved critical findings involving product quality, data integrity, sterility assurance, or regulatory compliance as potential disqualifiers regardless of the total weighted score.

Comparing Capacity, Costs, Timelines, and Supply Chain Strength

Choosing a biopharma manufacturing partner requires more than checking available equipment. Compare usable capacity, not headline capacity. A large bioreactor may still lack qualified operators, validated rooms, or flexible production slots. Ask how the facility manages technology transfer, scale changes, and unexpected deviations. Timelines should include engineering batches, quality testing, documentation, and batch release. Fast promises can hide missing steps.

Costs also need careful review. Request a detailed estimate covering materials, method validation, storage, shipping, testing, and change controls. The lowest quotation may become expensive after repeated revisions. Supply chain strength matters just as much. Examine raw material sourcing, supplier qualification, cold-chain monitoring, and backup transportation routes. One weak link can delay delivery by weeks. In practice, no continuity plan is perfect. That is worth admitting early.

Tips: Build a comparison table with capacity, total cost, timeline, and supply risks. Ask for evidence from comparable projects, not general assurances. Review deviation records and service-level expectations. Leave contingency time. I have seen schedules fail because teams trusted optimistic assumptions, including their own.

Selecting the Partner Through Due Diligence and Risk Review

How to Choose a Biopharma Manufacturing Partner?

Selecting a partner requires more than reviewing facility photos. Examine inspection history, deviation trends, batch-release records, and staffing stability. The FDA’s Drug Shortages Task Force report found that quality problems contributed to 62% of drug shortages. This makes quality-system due diligence a supply decision, not just a compliance exercise. Ask for evidence, not polished presentations.

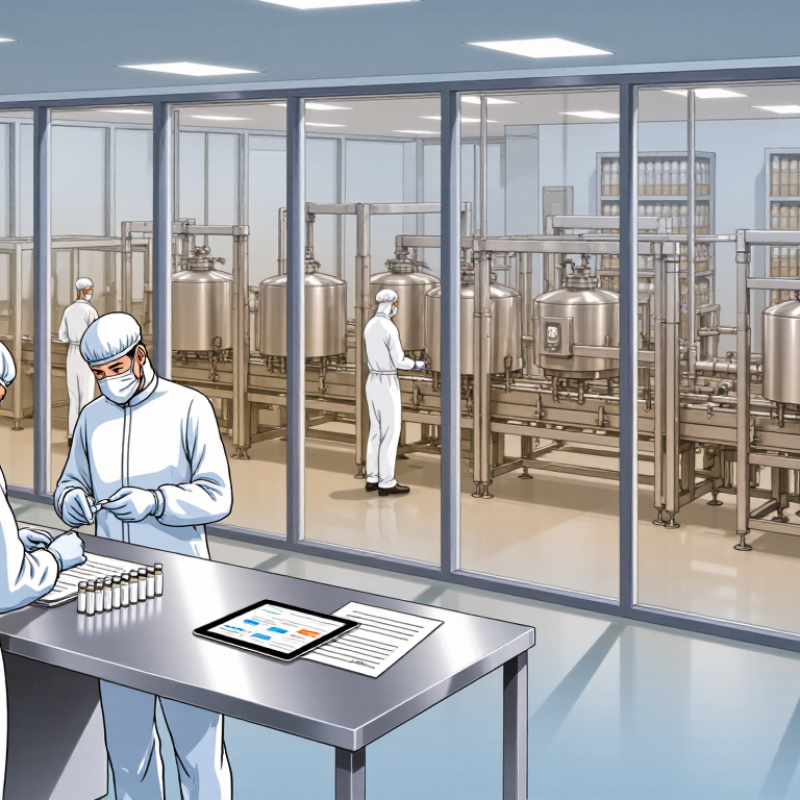
Review data integrity controls, change-control timelines, and corrective-action effectiveness. Walk through the site, if possible. Observe material flows, gowning practices, and environmental monitoring discussions. Compare the partner’s capacity claims with equipment logs and validated process ranges. The 2024 BioPlan Annual Report highlights continuing concerns around capacity, technology transfer, and operational reliability across biopharmaceutical manufacturing. A risk review should test these weak points before signing.

Tips:

Request three years of inspection and deviation summaries. Interview quality leaders without commercial staff present. Confirm backup suppliers, critical utilities, and business-continuity drills. Build a risk register with owners and deadlines. Do not accept “zero risk.” That answer deserves more questions. A smaller warning may reveal a stronger culture than a perfect-looking dashboard. Some assumptions will be wrong. Revisit them after the technical transfer, because early confidence can hide practical gaps.